
 Submit RFI/RFP

 Request Consultation
Introduction
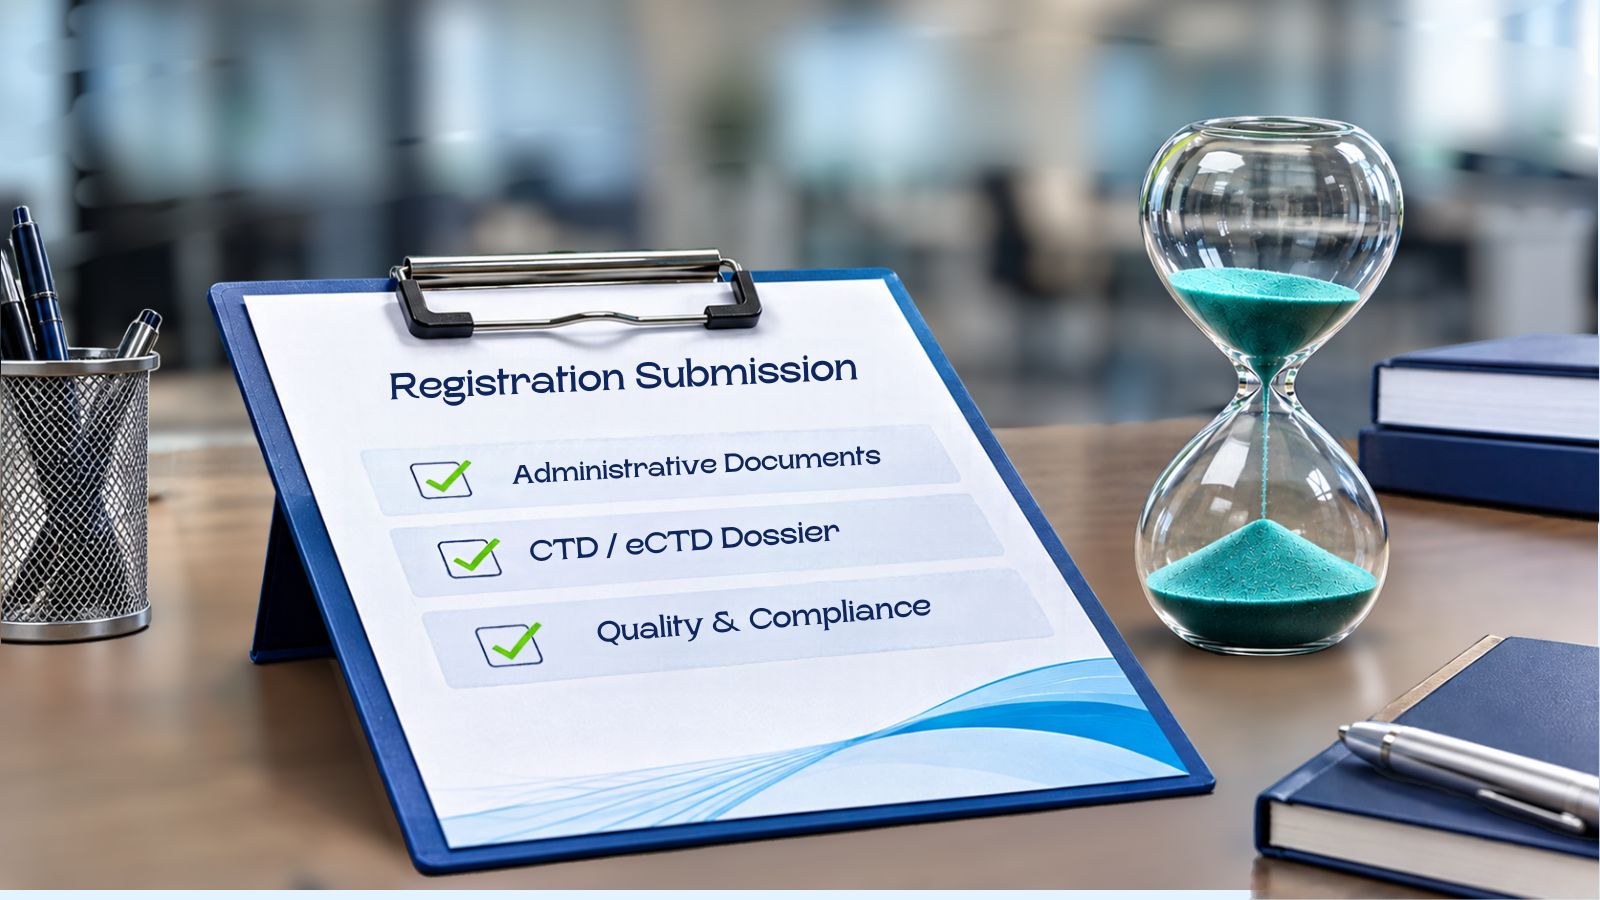
Pharmaceutical product registration is a critical step in bringing medicinal products to the market. It ensures that medicines meet the required standards of quality, safety, and efficacy before they become available to patients. However, obtaining marketing authorization is not always a straightforward process. Despite significant preparation and coordination, registration applications can experience unexpected delays due to incomplete documentation, inconsistencies in submitted data, regulatory non-compliance, or inadequate communication between stakeholders.
A delayed registration application does not necessarily indicate a problem with the medicinal product itself. In many cases, the delay results from avoidable gaps in the submission process, insufficient document preparation, or a lack of understanding of the applicable regulatory requirements.
Regulatory authorities assess applications against specific national and regional requirements, and even minor discrepancies can result in validation deficiencies, requests for additional information, or significant extensions to the review timeline.
Understanding the common causes of these delays is therefore essential for pharmaceutical companies, Marketing Authorization Holders (MAHs), regulatory affairs professionals, and other stakeholders involved in the registration process.
This article explores the most frequent gaps in pharmaceutical registration submissions, explains their potential impact, and highlights practical strategies to ensure a smoother and more efficient regulatory pathway.
1. Incomplete or Incorrect Administrative Documentation
One of the most common reasons for registration delays is the submission of incomplete, outdated, or incorrectly prepared administrative documents.
Although administrative documentation may appear straightforward compared with the scientific and technical sections of a registration dossier, it plays a fundamental role in application validation.
Typical administrative gaps include:
- Missing or expired manufacturing licenses.
- Outdated Good Manufacturing Practice (GMP) certificates.
- Missing or improperly legalized Power of Attorney (POA) documents.
- Incomplete Certificates of Pharmaceutical Product (CPPs), where required.
- Inconsistencies in the Marketing Authorization Holder’s legal name or address.
- Missing company registration documents or commercial registry extracts.
- Incorrect application forms or missing signatures.
- Documents that do not meet the authority’s language, translation, or legalization requirements.
Roles and Responsibilities
The Regulatory Affairs (RA) team is responsible for identifying the applicable administrative requirements, preparing a country-specific checklist, and verifying that all supporting documents are valid at the time of submission.
The MAH and manufacturer must provide accurate and up-to-date corporate and product-related documents. In addition, local regulatory representatives or authorized agents may be responsible for coordinating document legalization, translation, and submission to the national authority.
How to Prevent Delays
Companies should establish a centralized document management system that includes document validity dates, renewal timelines, responsible persons, and country-specific requirements.
A pre-submission administrative review should be conducted to confirm that all documents are complete, consistent, valid, and compliant with the relevant authority’s requirements.
2. Technical Documentation Gaps in the CTD/eCTD Dossier
The Common Technical Document (CTD) is the internationally recognized format used for organizing the scientific and technical information required for pharmaceutical registration.
The dossier is generally structured into five modules:
- Module 1: Regional administrative information and prescribing information.
- Module 2: CTD summaries and overviews.
- Module 3: Quality documentation.
- Module 4: Non-clinical study reports.
- Module 5: Clinical study reports.
Depending on the product type, application pathway, and applicable regulatory requirements, certain modules or documents may not be applicable.
A frequent source of registration delays is the submission of incomplete or inconsistent information across these modules.
Examples include:
- Missing sections or incomplete technical reports.
- Differences in product specifications between Module 2 and Module 3.
- Inconsistencies in the manufacturing process description.
- Missing validation data or insufficient supporting evidence.
- Inadequate justification of specifications.
- Missing or incomplete stability study reports.
- Inconsistencies between the submitted dossier and the proposed product information.
For eCTD submissions, additional issues may arise from incorrect document placement, technical validation errors, hyperlink problems, or failure to comply with the authority’s electronic submission specifications.
Roles and Responsibilities
The Regulatory Affairs team coordinates dossier preparation and ensures that the submission follows the applicable CTD/eCTD structure.
The Quality Assurance (QA), Quality Control (QC), Manufacturing, and Research and Development (R&D) teams are responsible for providing accurate and scientifically supported technical information.
The RA team must also perform a cross-functional consistency review to ensure that information provided by different departments is aligned.
How to Prevent Delays
A structured dossier gap assessment should be performed before submission.
Each module should undergo a technical review, followed by a cross-module consistency check. For electronic submissions, technical validation should be completed using the applicable validation criteria before the dossier is transmitted to the regulatory authority.
3. Quality Documentation and Manufacturing Compliance Issues
Product quality is one of the most important elements assessed during pharmaceutical registration.
Regulatory authorities need sufficient evidence that the medicinal product can be consistently manufactured according to established quality standards.
Registration applications may be delayed when the submitted quality documentation is incomplete, inconsistent, or insufficiently supported.
Common gaps include:
- Inadequate descriptions of the manufacturing process.
- Missing process validation reports or commitments, where applicable.
- Incomplete analytical method validation.
- Insufficient stability data to support the proposed shelf life.
- Inconsistencies between the manufacturing site details and GMP documentation.
- Inadequate justification of finished product specifications.
- Missing information regarding raw materials, excipients, or packaging materials.
- Unresolved manufacturing site compliance concerns.
For certain products, regulatory authorities may also request additional information regarding bioequivalence, comparability, or other product-specific quality requirements.
Roles and Responsibilities
The manufacturer is responsible for generating and maintaining accurate quality documentation and ensuring compliance with applicable Good Manufacturing Practice (GMP) requirements.
QA oversees quality systems and ensures that the necessary quality evidence is available and properly controlled.
QC provides analytical data, specifications, and validation information.
The RA team reviews the available documentation against regulatory requirements and ensures that identified gaps are addressed before submission.
How to Prevent Delays
Early coordination between RA, QA, QC, and manufacturing teams is essential.
A quality documentation checklist should be developed for each product, covering manufacturing information, specifications, analytical validation, stability data, and applicable GMP requirements.
Any identified gaps should be addressed before the application reaches the submission stage.
4. Stability Data and Shelf-Life Justification Deficiencies
Stability data are essential to demonstrate that a medicinal product maintains its quality throughout its proposed shelf life under specified storage conditions.
Insufficient or inappropriate stability documentation can lead to regulatory questions and requests for additional studies.
Common deficiencies include:
- Insufficient stability data to support the proposed shelf life.
- Inappropriate storage conditions or study design.
- Missing long-term or accelerated stability results.
- Inadequate justification of the proposed shelf life.
- Differences between the stability data and the proposed storage conditions stated in the product information.
- Missing stability data for the proposed packaging configuration.
- Failure to address significant changes or out-of-specification results.
Roles and Responsibilities
The manufacturer and stability teams are responsible for conducting stability studies in accordance with applicable guidelines and maintaining scientifically sound supporting data.
The RA team must verify that the submitted stability package supports the proposed shelf life, storage conditions, and packaging configuration and complies with the requirements of the target market.
How to Prevent Delays
Stability requirements should be assessed early during the registration planning stage.
A product-specific stability review should be performed to identify any missing data, inconsistencies, or additional studies that may be required by the target regulatory authority.
5. Inconsistencies in Product Information and Labeling
Product information is another important area that can lead to registration deficiencies.
Regulatory authorities assess proposed labeling and prescribing information to ensure that the information provided to healthcare professionals and patients is accurate, consistent, and compliant with local requirements.
Common gaps include:
- Differences between the SmPC, PIL, and packaging artwork.
- Incorrect product strength, dosage form, or pack size.
- Inconsistent manufacturing or MAH details.
- Missing mandatory warnings or safety statements.
- Non-compliance with local language requirements.
- Incorrect or missing storage conditions.
- Failure to include authority-specific labeling requirements.
- Differences between approved reference information and the submitted local product information.
These inconsistencies may require multiple rounds of clarification and artwork revisions.
Roles and Responsibilities
The RA team is responsible for reviewing product information against the applicable national requirements and ensuring consistency with the approved reference information, where relevant.
The Medical and Pharmacovigilance departments contribute to the accuracy of safety-related information, including contraindications, warnings, precautions, and adverse reaction information.
The Artwork and Packaging teams are responsible for implementing the approved content accurately in the final packaging materials.
How to Prevent Delays
A comprehensive labeling review should be performed before submission.
A cross-functional review involving RA, Medical, PV, and Artwork teams can help identify discrepancies before they become regulatory findings.
A controlled version-management process should also be implemented to ensure that only the latest approved documents are used.
6. Failure to Meet Country-Specific Regulatory Requirements
Although international guidelines provide a common framework for pharmaceutical registration, regulatory requirements are not identical across all countries.
Each national regulatory authority may have specific requirements regarding administrative documents, dossier formats, local representatives, product information, testing, legalization, and submission procedures.
For example, a document accepted by one authority may require additional certification, legalization, or translation in another country.
Other differences may include:
- National application forms and templates.
- Local MAH or authorized representative requirements.
- Country-specific labeling requirements.
- Local testing or sample submission requirements.
- Additional quality or clinical documentation.
- Specific requirements for reference countries or CPPs.
- National electronic submission platforms and procedures.
Submitting a dossier based solely on general international requirements without assessing the target country’s specific expectations can result in validation deficiencies or regulatory queries.
Roles and Responsibilities
The Regulatory Affairs team must identify and interpret the applicable country-specific requirements before initiating the registration process.
Local regulatory representatives are valuable sources of information regarding national submission procedures, authority expectations, and changes in regulatory requirements.
The MAH must ensure that the registration strategy and available documentation support the requirements of each intended market.
How to Prevent Delays
A country-specific regulatory assessment should be conducted before dossier preparation.
Companies should maintain updated regulatory intelligence trackers and establish communication channels with local representatives to monitor changes in national requirements.
A single global dossier should not automatically be assumed to be suitable for every market without the necessary regional adaptations.
7. Delayed Responses to Regulatory Authority Queries
Receiving questions or deficiency letters from regulatory authorities is a normal part of the registration process.
However, delays in responding to these queries can significantly extend the overall review timeline.
Regulatory authorities may request:
- Additional scientific or technical justification.
- Clarification of manufacturing procedures.
- Updated stability data.
- Revised product information or artwork.
- Additional safety or efficacy information.
- Clarification of administrative documentation.
- Corrective actions to address identified deficiencies.
Problems arise when responses are delayed, incomplete, poorly supported, or fail to address the authority’s actual concerns.
Roles and Responsibilities
The Regulatory Affairs team typically acts as the central coordinator for regulatory queries.
It is responsible for receiving and interpreting the authority’s comments, assigning questions to the relevant departments, consolidating responses, and ensuring submission within the established deadline.
Other departments must provide accurate, evidence-based responses within the agreed timelines.
The MAH or manufacturer remains responsible for ensuring that the final response is scientifically and technically justified.
How to Prevent Delays
An effective regulatory query management process should include:
- Recording each authority question in a centralized query tracker.
- Assigning a responsible department and response owner.
- Establishing internal deadlines ahead of the authority’s deadline.
- Reviewing supporting evidence and references.
- Conducting a final RA quality check before submission.
- Tracking the authority’s feedback and any subsequent actions.
A clear responsibility matrix can improve accountability and prevent unnecessary communication delays.
8. Poor Cross-Functional Coordination and Project Management
Pharmaceutical registration is not the responsibility of the Regulatory Affairs department alone. It is a multidisciplinary process involving several departments and external stakeholders.
A registration application may be delayed even when all the required information exists, simply because the relevant documents are not collected, reviewed, or approved on time.
For example, RA may be waiting for updated GMP documentation from QA, stability reports from the manufacturer, revised artwork from the packaging team, or safety-related information from PV.
Without clear communication and defined responsibilities, these dependencies can become significant bottlenecks.
Roles and Responsibilities
- Regulatory Affairs: Leads the regulatory strategy, dossier preparation, submission, and authority communication.
- Quality Assurance: Provides quality system documentation and relevant GMP evidence.
- Quality Control: Supplies analytical specifications, validation data, and quality results.
- Manufacturing: Provides manufacturing process information and supporting technical documentation.
- Research and Development: Supports formulation, development, and product characterization information.
- Medical Affairs: Reviews relevant clinical and medical information.
- Pharmacovigilance: Supports safety-related product information and applicable pharmacovigilance documentation.
- Artwork and Packaging: Ensures that packaging materials reflect approved product information.
- Project Management: Coordinates timelines, responsibilities, deliverables, dependencies, and escalation of delays.
How to Prevent Delays
A well-defined registration project plan should be established from the beginning.
It should include:
- Clear deliverables and responsible owners.
- Internal document submission deadlines.
- Regulatory submission milestones.
- Dependencies between departments.
- Document review and approval timelines.
- Risk identification and escalation procedures.
Regular progress meetings and centralized trackers can help identify bottlenecks before they affect the planned submission date.
9. Building an Effective Pre-Submission Quality Review
Preventing registration delays requires more than collecting the necessary documents. It requires a systematic process to verify that the application is complete, accurate, consistent, and aligned with the target authority’s requirements.
A pre-submission quality review should cover three essential areas:
1. Administrative Readiness
Confirm that all administrative documents are available, valid, signed, and compliant with country-specific requirements.
2. Technical and Scientific Readiness
Verify that the CTD/eCTD dossier is complete, scientifically supported, internally consistent, and technically validated where applicable.
3. Regulatory and Operational Readiness
Confirm that the submission strategy, country-specific requirements, timelines, responsibilities, and internal approvals are clearly established.
A formal pre-submission checklist, supported by a documented review and approval process, can significantly reduce avoidable deficiencies.
It is also important to recognize that not every regulatory query can be prevented. Authorities may request clarification or additional evidence as part of their scientific assessment. The objective is therefore to minimize avoidable deficiencies and ensure that any necessary responses are handled efficiently.
Baupharma Regulatory Submission Support
Baupharma supports pharmaceutical companies through Regulatory Affairs & Market Entry, offering regulatory dossier preparation, CTD module development, eCTD and NeeS publishing, country-specific adaptation, and health authority query management. Through structured regulatory assessments and coordinated submission support, these services help companies address documentation gaps, improve submission quality, and meet the applicable requirements of their target markets.
Key Takeaways
Pharmaceutical registration delays are often the result of preventable documentation gaps, technical inconsistencies, regulatory non-compliance, or ineffective coordination between stakeholders.
The most important lessons are:
- Preparation starts before submission: Conduct a comprehensive gap assessment before initiating the registration process.
- Administrative accuracy matters: Ensure that all supporting documents are valid, complete, and properly legalized where required.
- Dossier consistency is essential: Information must be aligned across all CTD modules and supporting documents.
- Quality documentation must be evidence-based: Manufacturing, analytical, and stability data should adequately support the proposed product.
- Country-specific requirements cannot be overlooked: Every target market requires an appropriate regulatory assessment.
- Timely communication makes a difference: Regulatory queries should be managed through clear responsibilities and structured response timelines.
- Cross-functional collaboration drives success: Effective coordination between RA, QA, QC, Manufacturing, Medical, PV, and Project Management is essential.
- Prevention is more efficient than correction: A robust pre-submission review can reduce avoidable deficiencies, rework, and registration delays.
Ultimately, successful pharmaceutical registration is not simply about submitting a dossier. It is about submitting the right information, in the right format, at the right time, while meeting the expectations of the relevant regulatory authority.
A well-prepared regulatory submission is an investment in efficiency, compliance, and ultimately, timely patient access to safe and effective medicines.



 Submit RFI/RFP